1
LEaP
Commands
Page 157
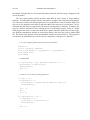
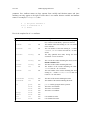
8.4.9. bond
bond atom1 atom2 [ order ]
ATOM
atom1
ATOM
atom2
STRING order
Create a bond between atom1 and atom2. Both of these ATOMs must be contained by the
same UNIT. By default, the bond will be a single bond. By specifying "-", "=", "#", or ":" as
the optional argument, order, the user can specify a single, double, triple, or aromatic bond,
respectively. Example:
bond trx.32.SG trx.35.SG
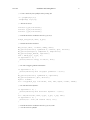
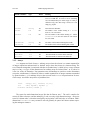
8.4.10. bondByDistance
bondByDistance container [ maxBond ]
CONT
container
NUMBER maxBond
Create single bonds between all ATOMs in container that are within maxBond angstroms of
each other. If maxBond is not specified then a default distance will be used. This command is
especially useful in building molecules. Example:
bondByDistance alkylChain
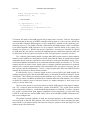
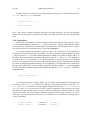
8.4.11. check
check unit [ parms ]
UNIT
unit
PARMSET parms
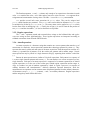
This command can be used to check the UNIT for internal inconsistencies that could cause
problems when performing calculations. This is a very useful command that should be used
before a UNIT is saved with saveAmberParm or its variants. Currently it checks for the following possible problems:
•
•
•
•
•
long bonds
short bonds
non-integral total charge of the UNIT.
missing force field atom types
close contacts (< 1.5 Å) between nonbonded ATOMs.
The user may collect any missing molecular mechanics parameters in a PARMSET for subsequent editing. In the following example, the alanine UNIT found in the amino acid library has