1
2.6. OPTIMIZATION OF THE MODEL
123
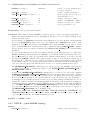
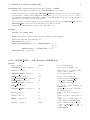
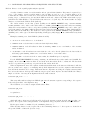
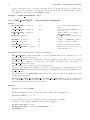
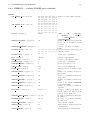
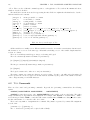
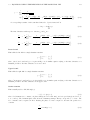
versus size n. The latter algorithm is used when the maximal difference in residue indices of the atoms in the
current dynamic restraints is larger than NLOGN USE or when the box size for this algorithm would have to
be larger than 8Å.
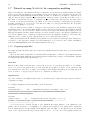
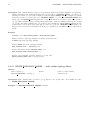
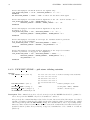
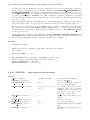
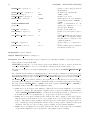
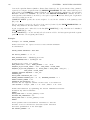
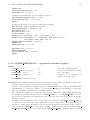
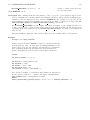
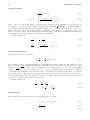
The molecular dynamics optimizer pretends that the natural logarithm of the molecular pdf is energy in
kcal/mole. MD TIME STEP is the time step in femtoseconds. TEMPERATURE is the temperature of the
system in degrees Kelvin. MAX ITERATIONS determines the number of MD steps. If MD RETURN is ’FINAL’
the last structure is returned as the MODEL. If MD RETURN is ’MINIMAL’ then the structure with the lowest
value of the objective function on the whole trajectory is returned as the MODEL. Rescaling of velocities
is done every EQUILIBRATION steps to match the specified temperature. Atomic shifts along one axis are
limited by CAP ATOM SHIFT. This value should be smaller than UPDATE DYNAMIC. If INIT VELOCITIES
= on, the velocity arrays are initialized, otherwise they are not. In that case, the final velocities from the
previous run are used as the initial velocities for the current run.
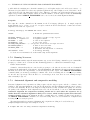
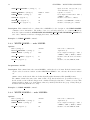
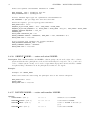
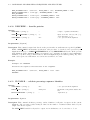
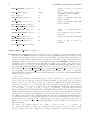
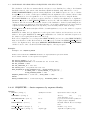
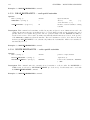
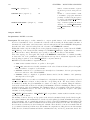
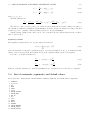
RAND SEED is the seed for the random number generator. It has to be between −2 and −50000. Its value
is changed after the return from the optimization routine.
MOLPDF contains the value of the objective function at the end of optimization.
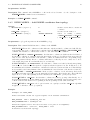
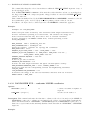
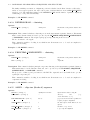
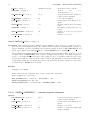
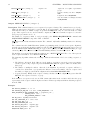
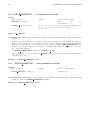
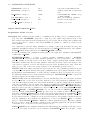
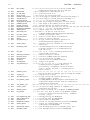
MODELLER STATUS is set to 1 if optimization is aborted because dynamic restraints could not be calculated
as a result of a system being too large. If MODELLER STATUS is equal or greater than STOP ON ERROR
the execution is stopped. Otherwise the execution returns back to the Top routine, exiting all optimization
routines immediately. The execution then continues as if nothing happened. It is up to the calling Top
routine to ensure that sensible action is taken; e.g., skipping the rest of modeling for the model that resulted
in an impossible function evaluation. This option is useful when calculating several independent models
and you do not want one bad model to abort the whole calculation. A probable reason for an interrupted
optimization is that it was far from convergence by the time the calculation of dynamic restraints was first
requested. Two possible solutions are: (1) optimize more thoroughly (i.e. slowly) and (2) use a different
contact pairs routine (SET NLOGN USE = 9999). MODELLER STATUS can be used in the Top routine to
exit from an optimization of a hopeless model and to continue with another model from a different initial
conformation.
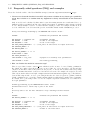
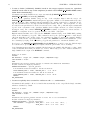
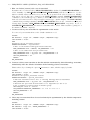
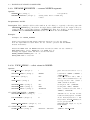
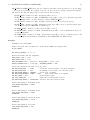
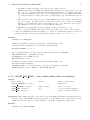
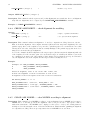
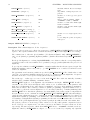
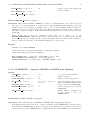
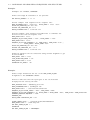
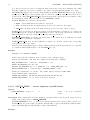
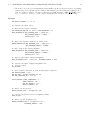
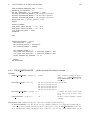
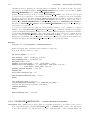
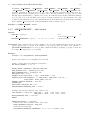
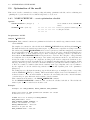
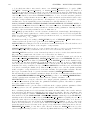
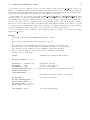
Example:
# Example for: OPTIMIZE, SWITCH_TRACE
# This will optimize stereochemistry of a given model, including
# non-bonded contacts.
READ_TOPOLOGY FILE = ’$(LIB)/top_heav.lib’
READ_PARAMETERS FILE = ’$(LIB)/par.lib’
READ_MODEL FILE = ’1fas’
SEQUENCE_TO_ALI ATOM_FILES = ’1fas’, ALIGN_CODES = ’1fas’
SEQUENCE_TO_ALI ADD_SEQUENCE = on, ATOM_FILES = ATOM_FILES ’1fas.ini’, ;
ALIGN_CODES = ALIGN_CODES ’1fas-ini’
GENERATE_TOPOLOGY SEQUENCE = ’1fas-ini’
TRANSFER_XYZ
BUILD_MODEL INITIALIZE_XYZ = off
WRITE_MODEL FILE = ’1fas.ini’
# Generate the restraints:
MAKE_RESTRAINTS RESTRAINT_TYPE = ’stereo’
WRITE_RESTRAINTS FILE = ’1fas.rsr’
ENERGY DYNAMIC_SPHERE = on
SWITCH_TRACE TRACE_OUTPUT = 1, FILE = ’1fas.trc’
OPTIMIZE OPTIMIZATION_METHOD = 1, MAX_ITERATIONS = 20
OPTIMIZE OPTIMIZATION_METHOD = 3, TEMPERATURE = 300, MAX_ITERATIONS = 50
OPTIMIZE OPTIMIZATION_METHOD = 1, MAX_ITERATIONS = 20
ENERGY