1
46
CHAPTER 2. MODELLER COMMANDS
#
#
#
#
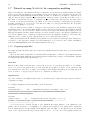
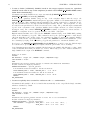
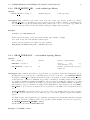
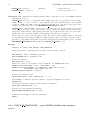
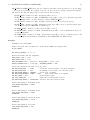
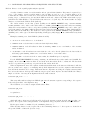
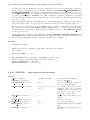
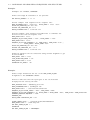
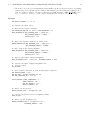
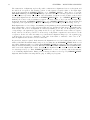
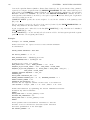
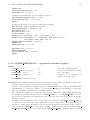
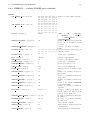
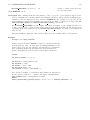
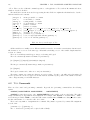
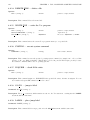
This will build a model for a given sequence by copying
coordinates from aligned templates. When the templates
have the same sequence as the target, this procedure ensures
that the new model corresponds to the MODELLER topology library.
READ_TOPOLOGY
FILE = ’$(LIB)/top_heav.lib’
READ_PARAMETERS FILE = ’$(LIB)/par.lib’
# Read the sequence and calculate its topology:
READ_ALIGNMENT FILE = ’toxin.ali’, ALIGN_CODES = ’2ctx’ ’1nbt’
MALIGN3D FIT = off
SET ADD_SEQUENCE = on
READ_ALIGNMENT FILE = ’toxin.ali’, ALIGN_CODES = ALIGN_CODES ’1fas’
GENERATE_TOPOLOGY SEQUENCE = ’1fas’
# Assign the average of the equivalent template coordinates to MODEL:
TRANSFER_XYZ
# Get the remaining undefined coordinates from internal coordinates:
BUILD_MODEL INITIALIZE_XYZ = off
# Write the final MODEL coordinates to a PDB file:
WRITE_MODEL FILE = ’1fas.ini’
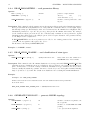
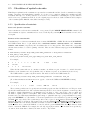
2.3.8
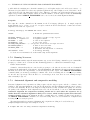
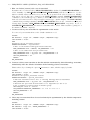
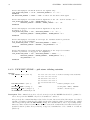
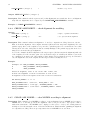
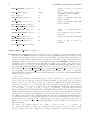
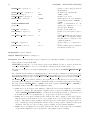
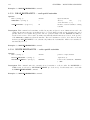
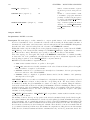
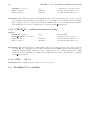
TRANSFER RES NUMB — residue numbers from MODEL2 to MODEL
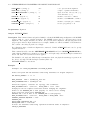
Options:
ALIGN CODES = hstring : 2i
’all’
MODEL2 code, MODEL code
Requirements: MODEL & MODEL2 [& alignment]
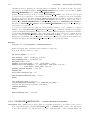
Description: This command transfers residue numbers and chain ids from MODEL2 to MODEL. It uses the
current alignment if present, otherwise a 1:1 correspondence is assumed. MODEL2 and MODEL must
correspond to the first and second protein in the alignment, respectively. The ALIGN CODES variable is used
only for output to the log file, not in the calculation. Both MODEL and MODEL2 must already be in
memory.
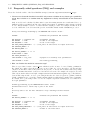
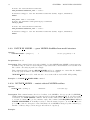
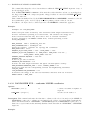
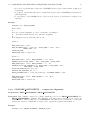
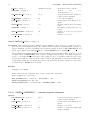
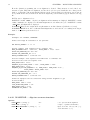
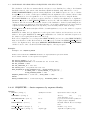
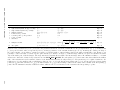
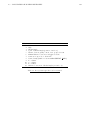
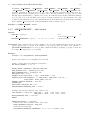
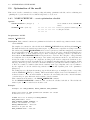
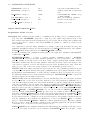
Example:
# Example for: TRANSFER_RES_NUMB
# This will transfer residue numbers and chain ids from model2 to model.
SET OUTPUT_CONTROL = 1 1 1 1 0
# Optionally, read an alignment for the transfer (otherwise 1:1 is assumed):
READ_ALIGNMENT FILE = ’toxin.ali’, ALIGN_CODES = ’2ctx’ ’1fas’
# Read the template and target models:
READ_MODEL2 FILE = ’2ctx’
READ_MODEL FILE = ’1fas’
# Transfer the residue and chain ids and write out the new MODEL:
TRANSFER_RES_NUMB
WRITE_MODEL FILE = ’1fas.ini’